Building and editing structures
The browser and desktop surfaces are an editor as well as a viewer. You can change a structure atom by atom, grow it from fragments, sketch one in vibe-sketch or build it from a name or a SMILES string, replicate a cell or cut a slab, and hand the result to a calculation: exported as an input script, or submitted to a queue without leaving the window.
Everything on this page runs in vibe-view open and vibe-view desktop.
The terminal and headless surfaces read structures; they do not edit them.
vibe-view owns detailed 3D structure work for calculations; vibe-sketch owns 2D chemical drawing and reaction figures for papers. Both retain input, drawing, basic optimisation and calculation workflows. See the shared scope decision, including vibe-sketch’s planned reference-geometry and FIRE cleanup.
Edit mode
Press e or the pencil icon. The Atom Editor card opens and the viewport starts taking clicks as edits:
Select an atom by clicking it; a selected atom carries an orange wireframe highlight. Click again to deselect, Delete to remove the selection.
Add an atom by clicking empty space. The element comes from the Element used for newly added atoms picker.
Change every selected atom’s element with the element picker in the card.
Undo / redo with Ctrl/Cmd+Z and Ctrl/Cmd+Shift+Z. Every edit is one undo step.
Bonds are recomputed after each edit with the same covalent-radius rule the viewer draws with, so connectivity does not change on the first edit, and a periodic structure keeps its bonds through the cell faces. The edited structure is drawn the way the unedited one is, in the representation and at the replication you chose. Edits compound in place: the geometry you export or submit is the geometry as edited, and the original archive on disk is not touched.
An edit replaces the atom list, so the file’s biomolecular metadata does not carry over to the edited geometry: residue and chain names, B-factors, and any producer-supplied residues, chains or secondary structure describe the original atoms and are dropped. The views built on that metadata (cartoon and backbone, colouring by chain, secondary structure or B-factor) have nothing to draw from until the archive is reloaded.
Edit mode and measure mode are mutually exclusive; Esc leaves either.
Drag atoms and selections
Select Structure, press e to open Atom Editor, then enable Drag atoms. Press an atom and drag it to move it. An atom outside the current selection becomes the new selection when you grab it.
Shift-click atoms to add them to, or remove them from, the selection. Drag any selected atom to translate all selected, unfrozen atoms together.
Select connected expands the selection through bonds, so a connected molecule or fragment can be moved as a group. Start by selecting one or more atoms in the desired groups. Connectivity is the same as in the viewer.
Select all selects every atom; Clear selection deselects everything.
Freeze Selected keeps those atoms fixed, including when a larger selection is dragged. Unfreeze All releases them.
The movement follows the screen plane through the atom you grabbed. Rotate the view and drag again to move along another direction in 3D. In drag mode, Alt-drag rotates the camera, the middle mouse button pans and the wheel zooms. Clicking empty space does not add an atom while Drag atoms is on; turn it off to return to the ordinary add/select editor.
Every drag is one Undo step, regardless of the number of intermediate positions shown. Redo reapplies the final coordinates. Releasing outside the viewport or losing focus finishes at the last received position. If Auto-optimize is enabled, it pauses during a drag and resumes on release. Turn it off when the moved coordinates should remain unchanged for export. For periodic structures, use replication 1 × 1 × 1 when dragging atoms.
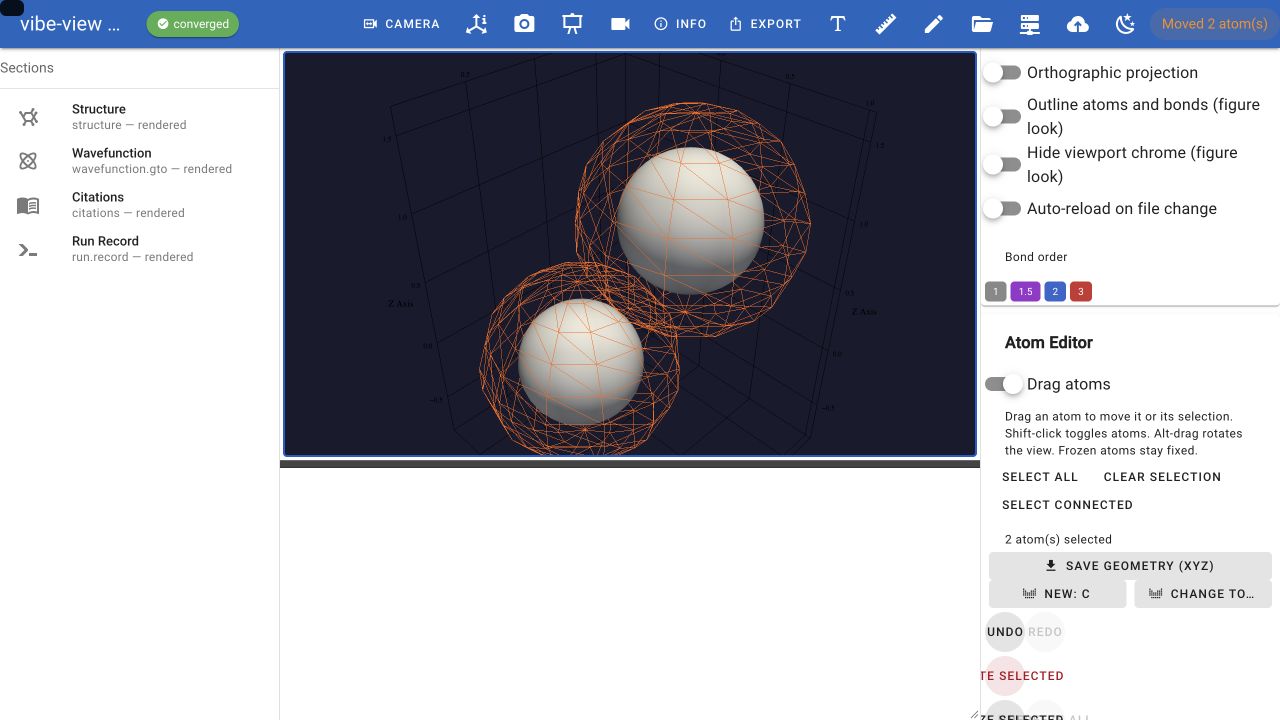
Save geometry (XYZ) and generated calculation inputs use the edited coordinates. Dragging never rewrites the original loaded result file.
Save coordinates and break symmetry
In Atom Editor, Save geometry (XYZ) downloads the current edited geometry as plain text: atom count, comment, then element and Cartesian coordinates in Å. The toolbar’s Export → XYZ does the same. The file can be opened in any text editor; it contains the edited coordinates, not the original coordinates from the loaded calculation.
To introduce a small random perturbation, set Maximum displacement (Å) (default 0.01, allowed range greater than zero through 0.1) and click Random displacement. Selected atoms move; if none are selected, all atoms move. Frozen atoms always stay fixed. Each Cartesian component is sampled independently within a cube chosen so that the total displacement of each atom never exceeds the specified maximum. Repeated clicks produce new perturbations. Each click is one Undo step; Redo restores exactly that perturbation.
Displacement switches Auto-optimize off, including any pending run, so the new coordinates remain available for export. Use it after pre-optimising if the calculation should start from a slightly asymmetric structure. This is a geometry perturbation tool, not a guarantee that a calculation’s linear-dependence warning will disappear.
Displace along a normal mode
You can displace along one mode or a combination of selected modes.
Open a calculation containing normal modes (for ORCA, include its .hess
file) and select Vibrations. Choose a Mode, then enter Signed peak
displacement (Å) and click Create displaced structure. For example,
0.10 and −0.10 produce opposite distortions. Nonzero values through
±1 Å are accepted; the default is 0.10 Å.
For a combination, choose two or more entries in Modes to displace. Each selected mode gets its own signed peak displacement (Å) field. For example, set one mode to +0.10 Å and another to −0.05 Å, then click Create displaced structure. Only the selected modes contribute; the animation’s Mode selection does not add another contribution. Clear Modes to displace to return to the single-mode workflow above.
The viewer returns to Structure with Atom Editor open. Use Save geometry (XYZ), prepare an ORCA input, or submit it through vq. Undo restores the geometry before the displacement; Redo restores the exact displaced coordinates. Animation and auto-optimisation stop when you apply it.
Each application uses the mode’s equilibrium geometry, replacing current coordinate edits in one undoable step. It never adds another displacement on top of the last one. To make both directions, save the positive geometry, return to Vibrations, enter the negative value and apply again.
For each mode, the longest atomic displacement vector has the absolute length you enter;
all other vectors scale by the same factor. The mode contributions are added:
R = R_equilibrium + Σₖ aₖ dₖ / maxᵢ |dₖᵢ|. The sum is not renormalized, so
its largest atomic movement can exceed an individual amplitude or cancel
through opposing contributions. Each coefficient must be nonzero and within
±1 Å; deselect a mode to omit it. Selections reset when switching sections
or calculations. This uses the stored Cartesian
mode, without further mass weighting, and is independent of the animation
amplitude or phase. Both real and imaginary modes can be used. The sign of
a mode vector is arbitrary, so “positive” refers to the direction stored in
that calculation. This is a geometry preparation tool, not an IRC trajectory
or a thermal amplitude.
The full mode moves the molecule regardless of atom selection. Unfreeze All first if atoms are frozen. The operation rejects zero/non-finite mode vectors, changed atom lists, and modes whose equilibrium geometry or associated structure does not match. Legacy modes with all-zero equilibrium coordinates use the archived structure, never its edited coordinates. Original frequencies, spectra and orbitals still describe the original calculation; obtain new properties with a new calculation.
The ORCA peroxide tutorial walks through both directions using a real frequency result. For several directions at once, follow Combine selected modes: viewer Modes 7 and 8 have signed peak displacements of +0.10 and −0.05 Å. The follow-up calculation submits that combined geometry to ORCA through vq and opens a new dataset with the resulting structure, orbitals and charges. Download either archive to inspect it without running ORCA yourself.
Fragments and hydrogens
The Fragment Library attaches a group to the last selected atom, one bond length (1.5 Å) out from it, on the side facing away from the molecule’s centre. With nothing selected, the group is set down beside the structure, 2.5 Å past its largest x coordinate, so it never lands inside the molecule:
Fragment |
Group |
|---|---|
|
methyl |
|
amino |
|
hydroxyl |
|
carboxyl |
|
phenyl |
|
aldehyde |
|
nitro |
|
cyano |
|
trifluoromethyl |
|
sulfonic acid |
Add Hydrogens saturates the open valences of the selected atoms with tetrahedral geometry where it applies. The same operation is available for a whole file from the command line:
vibe-view h-add structure.qvf -o structure_h.qvf
Sketching a molecule in vibe-sketch
vibe-sketch is the family’s 2D structure and reaction sketcher. When it is installed beside the viewer, Sketch in vibe-sketch at the top of the Build molecule dialog opens it in your browser. The command palette (Ctrl/Cmd+K) has the same action, as Sketch a molecule in vibe-sketch. Draw the molecule, then choose Send to vibe-view in the sketcher. The structure opens here as a new file and the status bar says how many atoms it has. Send again after an edit and the new version opens as another file, so you can compare it with the earlier one.
The sketcher runs on this machine at http://127.0.0.1:8766/, or the next
free port, and keeps running while the viewer is open. Clicking Sketch
again brings its page back rather than starting a second copy. Closing the
viewer also stops the sketcher, so save a drawing you want to keep in
vibe-sketch first.
vibe-sketch is the optional [sketch] extra. It is published on no package
index yet, so install it from a vibe-sketch checkout first. The extra is
then satisfied by what is already there:
python -m pip install <path-to-vibe-sketch>
python -m pip install -e '.[sketch]'   # from the vibe-view checkout
vibe-sketch needs Python 3.12 or newer. Without it the Sketch button stays
in the dialog, and clicking it puts the install command in the status bar.
vibe-view doctor lists the same command under sketch.
The hand-off between the two programs
This is the contract vibe-sketch implements for Send to vibe-view
(vibe-view #77, vibe-sketch #1). It lives in vibeview.sketch:
vibe-view starts
python -m vibesketch serve --host 127.0.0.1 --port PORT --openwith its own interpreter. The environment variableVIBEVIEW_SKETCH_HANDOFFnames a private directory created for this sketcher. When the variable is set, the sketcher knows a viewer is waiting.Send to vibe-view writes the structure as a
.qvfor.xyzfile, then writes a record into that directory: a JSON object{"schema": "vibe-view-sketch-handoff/1", "path": "<absolute path>"}. The record is written under a name that does not end in.jsonand then renamed to*.json, so the viewer never reads half of it.About once a second, vibe-view reads new records in name order and deletes each one after reading it. It reads the structure into memory, so a later send that overwrites the same file does not change what is already open. A record that breaks the contract, or names a file that cannot be read, is reported in the status bar and opens nothing.
When the viewer exits, it stops the sketcher and removes the directory. A missing directory tells the sketcher that the viewer is gone.
Building from a name or a SMILES string
Below the Sketch button, the Build molecule dialog takes either:
A molecule name. Geometries come from the curated structure database of the
vibeqc_namingpackage, which ships with vibe-qc. A standalone viewer install does not have it, so the name field is left out of the dialog. The toolbar button still opens the dialog when vibe-sketch is installed.A SMILES string. Needs the
[smiles]extra (RDKit). The geometry is an RDKit ETKDG embedding with no force-field cleanup: a chemically sensible starting point, not an optimised structure.
The Structure Library card remembers what you built, and lets you pin favourites to reuse. It shows a thumbnail of the molecule you pick, and each pinned and recent chip carries a small picture of its molecule. The pictures are drawn in the background from the library’s geometry once a name is picked, pinned or used, so they appear a moment after the name.
From Python the SMILES route is a function that returns an in-memory archive, so it can be handed straight to the viewer:
from vibeview import launch_qvf
from vibeview.converters import smiles_to_qvf

launch_qvf(smiles_to_qvf("c1ccccc1"))                 # benzene
launch_qvf(smiles_to_qvf("CCO", add_hydrogens=False))
There is no vibe-view subcommand for SMILES and no SMILES file format the
open dialog accepts; the dialog and the function are the two routes.
Live optimisation while you build
Auto-optimize, in the Atom Editor card, relaxes the sketch in the background after every pause in editing and streams each optimiser step into the viewport, so the atoms settle toward a relaxed geometry as you work. The status line shows the step, energy and maximum gradient, then relaxed in N steps. Editing again mid-relax cancels and reschedules, and each relax run is one undo entry.
This needs vibe-qc importable in the viewer’s environment: the default
engine is its MSINDO semi-empirical model, and when its [mace] extra is
installed an Engine picker offers the MACE foundation potential as well.
Without vibe-qc the switch snaps back off with the reason and the editor
works exactly as before. Structures beyond 80 atoms are skipped to keep the
background evaluations interactive.
Selected atoms can be frozen so a relaxation moves everything but them.
The Optimizer selector offers L-BFGS (default) and FIRE (damped MD). FIRE uses forces from the selected engine and adaptive, damped motion to relax the geometry, following Bitzek et al., Phys. Rev. Lett. 97, 170201 (2006). It does not need another package beyond the selected energy/force engine. Its timestep is a numerical relaxation parameter; this is not a finite-temperature MD trajectory. FIRE allows up to 200 steps, limits each atom’s move to 0.1 Å per step, and uses the same maximum Cartesian gradient tolerance as L-BFGS (0.002 Ha/bohr). If the step limit is reached, the status explicitly says not converged. Changing the optimiser while Auto-optimize is on starts a fresh run from the current geometry. Undo restores the geometry before that run.
For a pre-optimisation after drawing: select the optimiser, turn on Auto-optimize, wait for relaxed in N steps, then turn it off before exporting. See the ORCA builder walkthrough for the complete sequence.
Periodic structures
Build Supercell replicates the cell Nx × Ny × Nz into a new structure;
the command-line equivalent is:
vibe-view supercell si.qvf --nx 2 --ny 2 --nz 2 -o si_2x2x2.qvf
The crystal-builder helpers behind it are importable for scripts: a
space-group search, conversion between a lattice matrix and cell parameters,
Miller-plane slab cutting with a vacuum gap, and supercell replication.
miller_slab works for any orientation of a nonsingular cell: it builds the (hkl)
surface cell from two lattice vectors that lie in the plane and one that steps
a single interplanar spacing along the normal, repeats the parent cell’s atoms
n_layers times along that step, and adds 2 * vacuum to the cell length
along the normal. The atoms are not shifted to center them in that padding,
so the gaps on either side need not be equal. The in-plane vectors and the
atoms stay in the parent crystal’s Cartesian frame, so the slab overlays the
bulk it was cut from. The determinant-one surface basis preserves the input
cell’s handedness. n_layers
counts repetitions of the cell, so a conventional fcc cell gives one atomic
plane per layer for (111) and two for (001); indices with a common factor,
such as (002), name the same plane family as their reduced form.
from vibeview.crystal_builder import (
    search_space_groups, cell_from_abc, abc_from_cell, miller_slab, replicate_cell,
)

search_space_groups("Fm-3m")                          # [{'number': 225, ...}]
cell = cell_from_abc(5.43, 5.43, 5.43, 90, 90, 90)
a, b, c, alpha, beta, gamma = abc_from_cell(cell)
new_cell, slab_atoms = miller_slab(cell, atoms, hkl=(1, 1, 1), n_layers=3, vacuum=15.0)
super_cell, super_atoms = replicate_cell(cell, atoms, nx=2, ny=2, nz=2)
Cell edges are only drawn along periodic axes, so a slab gets its in-plane parallelogram and never a box around the vacuum.
From structure to calculation
The Calculation Parameters panel
Select Calculation engine → ORCA for basic molecular ORCA preparation,
input preview, .inp download and vq submission. The ORCA walkthrough
shows each step. The settings below describe the vibe-qc engine.
With a structure active, the right drawer shows Calculation Parameters: method (RHF, UHF, RKS, UKS, RMP2, UMP2), functional (PBE, PBE0, B3LYP, BLYP, BP86, TPSS, M06-2X, ωB97X-D, CAM-B3LYP, LDA, r²SCAN, HSE06), basis set (STO-3G through aug-cc-pVTZ and the def2 family), charge, multiplicity, and the calculation type: single point, geometry optimisation, frequencies, all three, or a periodic single point for a periodic structure. Charge and multiplicity start from the structure on show when the file declares them: the structure’s own chemical identity first, then the archive’s provenance when it holds that one structure only. A vibe-qc result, a vibe-sketch species, an ion built from SMILES, or an XYZ, SDF or Gaussian file stating them does. Otherwise they keep their current values. Showing another structure of the archive seeds them again from that structure.
Export vibe-qc input (.py) writes a vibe-qc input script with those
settings and the geometry as edited. Charge and multiplicity go on the
Molecule or PeriodicSystem, where vibe-qc reads them; the runner call
carries the method, basis and outputs. The same generator is behind
vibe-view export FILE -f py, which additionally reuses the archive’s own
provenance (and the structure’s own charge and multiplicity, when it states
them) so the regenerated script reproduces the original calculation,
and carries the lattice so a periodic file exports a periodic system rather
than silently degrading to a molecule.
The panel’s settings are also what a queue submission uses.
Submitting to the queue
Submit to vq cluster sends the current structure to a
vibe-queue daemon with the
parameters above and opens the Job Manager so the job appears immediately.
That needs the [queue] extra and vibe-queue itself; see
Queue integration for the install and Jobs and live
results for what happens after you press the button.
Reading an input script back in
The reverse direction works too. Open a vibe-qc Python input directly:
vibe-view open input.py
The parser walks the file’s syntax tree without executing it and extracts the structure and the run parameters, so the structure appears in the viewport and the Calculation Parameters panel is pre-filled from the script. It understands vibe-qc’s input syntax and the input library’s builder functions; it is not a general Python interpreter, and arbitrary code in an input resolves to nothing rather than running.
What the file on disk sees
None of this writes to the archive you opened. In an archive with several structures, an edit belongs to the structure it was made on and stays with it while you look at another; exports and submissions take the structure on show. To keep an edited structure:
Export it from the toolbar as XYZ, CIF, CML or a Python input, or as a mesh or scene; see Figures without a display for the formats.
Save a session to keep views of it; see Bookmarks and sessions.
Submit it, in which case the settled result comes back as its own archive.